#include <molecule.h>
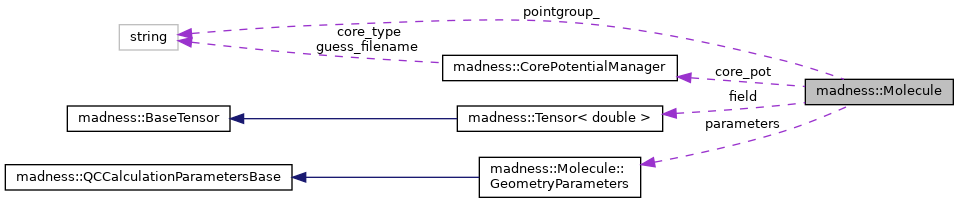
Classes | |
| struct | apply_c2 |
| struct | apply_inverse |
| struct | apply_sigma |
| struct | GeometryParameters |
Public Member Functions | |
| Molecule () | |
| Makes a molecule with zero atoms. | |
| Molecule (std::vector< Atom > atoms, double eprec, CorePotentialManager core_pot={}, madness::Tensor< double > field=madness::Tensor< double >(3L)) | |
| makes a molecule from a list of atoms | |
| Molecule (World &world, const commandlineparser &parser) | |
makes a molecule using contents of parser | |
| void | add_atom (double x, double y, double z, double q, int atn) |
| void | add_atom (double x, double y, double z, double q, int atn, bool psat) |
| double | atomic_attraction_potential (int iatom, double x, double y, double z) const |
| nuclear attraction potential for a specific atom in the molecule | |
| double | bounding_cube () const |
| Returns the half width of the bounding cube. | |
| void | center () |
| Moves the center of nuclear charge to the origin. | |
| Tensor< double > | center_of_mass () const |
| compute the center of mass | |
| double | core_derivative (int atom, int axis, unsigned int core, int m, double x, double y, double z) const |
| double | core_eval (int atom, unsigned int core, int m, double x, double y, double z) const |
| double | core_potential_derivative (int atom, int axis, double x, double y, double z) const |
| std::vector< std::string > | cubefile_header (const Vector< double, 3 > offset=Vector< double, 3 >(0.0)) const |
| print out a Gaussian cubefile header | |
| void | fix_phase () |
| rotates the molecule and the external field | |
| void | from_json (const json &mol_json) |
| madness::Tensor< double > | get_all_coords () const |
| std::vector< madness::Vector< double, 3 > > | get_all_coords_vec () const |
| const Atom & | get_atom (unsigned int i) const |
| unsigned int | get_atom_charge (unsigned int i) const |
| unsigned int | get_atomic_number (unsigned int i) const |
| const std::vector< Atom > & | get_atoms () const |
| double | get_core_bc (unsigned int atn, unsigned int c) const |
| unsigned int | get_core_l (unsigned int atn, unsigned int c) const |
| double | get_eprec () const |
| std::string | get_pointgroup () const |
| bool | get_pseudo_atom (unsigned int i) const |
| std::vector< double > | get_rcut () const |
| void | get_structure () |
| std::string | guess_file () const |
| hashT | hash () const |
| void | insert_symbols_and_geometry (json &mol_json) const |
| double | inter_atomic_distance (unsigned int i, unsigned int j) const |
| bool | is_potential_defined (unsigned int atn) const |
| bool | is_potential_defined_atom (int i) const |
| Tensor< double > | massweights () const |
| compute the mass-weighting matrix for the hessian | |
| double | molecular_core_potential (double x, double y, double z) const |
| Tensor< double > | moment_of_inertia () const |
| unsigned int | n_core_orb (unsigned int atn) const |
| unsigned int | n_core_orb_all () const |
| size_t | natom () const |
| double | nuclear_attraction_potential (double x, double y, double z) const |
| nuclear attraction potential for the whole molecule | |
| double | nuclear_attraction_potential_derivative (int atom, int axis, double x, double y, double z) const |
| double | nuclear_attraction_potential_second_derivative (int atom, int iaxis, int jaxis, double x, double y, double z) const |
| the second derivative of the (smoothed) nuclear potential Z/r | |
| double | nuclear_charge_density (double x, double y, double z, double rscale=1.) const |
| double | nuclear_dipole (int axis) const |
| compute the dipole moment of the nuclei | |
| Tensor< double > | nuclear_dipole_derivative (const int atom, const int axis) const |
| compute the derivative of the nuclear dipole wrt a nuclear displacement | |
| double | nuclear_repulsion_derivative (size_t iatom, int axis) const |
| double | nuclear_repulsion_energy () const |
| Tensor< double > | nuclear_repulsion_hessian () const |
| return the hessian matrix of the second derivatives d^2/dxdy V | |
| double | nuclear_repulsion_second_derivative (int iatom, int jatom, int iaxis, int jaxis) const |
| compute the nuclear-nuclear contribution to the second derivatives | |
| bool | operator== (const Molecule &other) const |
| void | orient (bool verbose=false) |
| Centers and orients the molecule in a standard manner. | |
| void | print (std::ostream &os=std::cout, const bool defined_only=false) const |
| print all parameters and the molecular geometry | |
| void | print_defined_only (std::ostream &os=std::cout) const |
| print user-defined parameters and the molecular geometry | |
| void | read (std::istream &f) |
| void | read_core_file (const std::string &filename) |
| void | read_file (const std::string &filename) |
| void | read_structure_from_library (const std::string &name) |
| void | read_xyz (const std::string filename) |
| void | rotate (const Tensor< double > &D) |
| template<typename Archive> | |
| void | serialize (Archive &ar) |
| void | set_all_coords (const madness::Tensor< double > &newcoords) |
| void | set_atom_charge (unsigned int i, double zeff) |
| void | set_atom_coords (unsigned int i, double x, double y, double z) |
| void | set_core_eprec (double value) |
| void | set_core_rcut (double value) |
| void | set_pseudo_atom (unsigned int i, bool psat) |
| void | set_rcut (double value) |
| double | smallest_length_scale () const |
| std::string | symmetrize_and_identify_point_group (const double symtol) |
| json | to_json () const |
| json | to_json_if_precedence (std::string const &precedence) const |
| double | total_nuclear_charge () const |
| void | translate (const Tensor< double > &translation) |
| translate the molecule | |
| void | update_rcut_with_eprec (double value) |
| updates rcuts with given eprec | |
Static Public Member Functions | |
| static std::string | get_structure_library_path () |
| static std::istream & | position_stream_in_library (std::ifstream &f, const std::string &name) |
| static void | print_parameters () |
Public Attributes | |
| std::vector< double > | atomic_radii |
| GeometryParameters | parameters |
Static Public Attributes | |
| static constexpr char const * | tag = "molecule" |
Private Member Functions | |
| template<typename opT> | |
| int | find_symmetry_equivalent_atom (int iatom, opT op, const double symtol) const |
| void | swapaxes (int ix, int iy) |
| template<typename opT> | |
| void | symmetrize_for_op (opT op, const double symtol) |
| bool | test_for_c2 (double xaxis, double yaxis, double zaxis, const double symtol) const |
| bool | test_for_inverse (const double symtol) const |
| template<typename opT> | |
| bool | test_for_op (opT op, const double symtol) const |
| bool | test_for_sigma (double xaxis, double yaxis, double zaxis, const double symtol) const |
Private Attributes | |
| std::vector< Atom > | atoms |
| CorePotentialManager | core_pot |
| madness::Tensor< double > | field |
| std::string | pointgroup_ = "c1" |
| std::vector< double > | rcut |
Constructor & Destructor Documentation
◆ Molecule() [1/3]
|
inline |
Makes a molecule with zero atoms.
References atoms, core_pot, field, and rcut.
Referenced by operator==().
◆ Molecule() [2/3]
| madness::Molecule::Molecule | ( | std::vector< Atom > | atoms, |
| double | eprec, | ||
| CorePotentialManager | core_pot = {}, | ||
| madness::Tensor< double > | field = madness::Tensor<double>(3L) ) |
makes a molecule from a list of atoms
References atomic_radii, atoms, core_pot, eprec, and field.
◆ Molecule() [3/3]
| madness::Molecule::Molecule | ( | World & | world, |
| const commandlineparser & | parser ) |
makes a molecule using contents of parser
References atoms, madness::WorldGopInterface::broadcast_serializable(), core_pot, field, get_structure(), madness::World::gop, MADNESS_CHECK, parameters, madness::World::rank(), rcut, and read_core_file().
Member Function Documentation
◆ add_atom() [1/2]
| void madness::Molecule::add_atom | ( | double | x, |
| double | y, | ||
| double | z, | ||
| double | q, | ||
| int | atn ) |
References atomic_radii, madness::constants::atomic_unit_of_length, atoms, c, madness::AtomicData::covalent_radius, e(), madness::get_atomic_data(), get_eprec(), q(), rcut, and madness::smoothing_parameter().
Referenced by from_json(), main(), read(), and read_xyz().
◆ add_atom() [2/2]
| void madness::Molecule::add_atom | ( | double | x, |
| double | y, | ||
| double | z, | ||
| double | q, | ||
| int | atn, | ||
| bool | psat ) |
◆ atomic_attraction_potential()
| double madness::Molecule::atomic_attraction_potential | ( | int | iatom, |
| double | x, | ||
| double | y, | ||
| double | z ) const |
nuclear attraction potential for a specific atom in the molecule
References atoms, madness::distance(), rcut, madness::smoothed_potential(), and madness::sum().
◆ bounding_cube()
| double madness::Molecule::bounding_cube | ( | ) | const |
◆ center()
| void madness::Molecule::center | ( | ) |
Moves the center of nuclear charge to the origin.
References atoms, and translate().
Referenced by orient().
◆ center_of_mass()
| Tensor< double > madness::Molecule::center_of_mass | ( | ) | const |
compute the center of mass
References get_atom(), madness::Atom::mass, natom(), madness::Atom::x, madness::Atom::y, and madness::Atom::z.
Referenced by madness::MolecularOptimizer::projector_external_dof(), and projector_external_dof().
◆ core_derivative()
| double madness::Molecule::core_derivative | ( | int | atom, |
| int | axis, | ||
| unsigned int | core, | ||
| int | m, | ||
| double | x, | ||
| double | y, | ||
| double | z ) const |
◆ core_eval()
| double madness::Molecule::core_eval | ( | int | atom, |
| unsigned int | core, | ||
| int | m, | ||
| double | x, | ||
| double | y, | ||
| double | z ) const |
◆ core_potential_derivative()
| double madness::Molecule::core_potential_derivative | ( | int | atom, |
| int | axis, | ||
| double | x, | ||
| double | y, | ||
| double | z ) const |
◆ cubefile_header()
| std::vector< std::string > madness::Molecule::cubefile_header | ( | const Vector< double, 3 > | offset = Vector<double, 3>(0.0) | ) | const |
print out a Gaussian cubefile header
- Parameters
-
[in] offset the offset to be subtracted from the coordinates
References get_atom(), madness::Atom::get_atomic_number(), madness::Atom::get_coords(), natom(), and offset.
◆ find_symmetry_equivalent_atom()
|
private |
References atoms, madness::distance(), and op().
Referenced by symmetrize_for_op(), and test_for_op().
◆ fix_phase()
| void madness::Molecule::fix_phase | ( | ) |
◆ from_json()
| void madness::Molecule::from_json | ( | const json & | mol_json | ) |
References add_atom(), madness::constants::atomic_unit_of_length, atoms, e(), orient(), parameters, q(), rcut, madness::scale(), and madness::symbol_to_atomic_number().
◆ get_all_coords()
| madness::Tensor< double > madness::Molecule::get_all_coords | ( | ) | const |
References c, get_atom(), natom(), madness::Atom::x, madness::Atom::y, and madness::Atom::z.
Referenced by madness::DispersionCorrection::compute(), madness::DispersionCorrection::hessian(), main(), run_all_calculations(), test_gradient(), test_hessian(), test_read_restartaodata(), and test_read_restartdata().
◆ get_all_coords_vec()
| std::vector< madness::Vector< double, 3 > > madness::Molecule::get_all_coords_vec | ( | ) | const |
References c, get_atom(), natom(), madness::Atom::x, madness::Atom::y, and madness::Atom::z.
◆ get_atom()
| const Atom & madness::Molecule::get_atom | ( | unsigned int | i | ) | const |
References atoms.
Referenced by center_of_mass(), madness::compare_geometry(), cubefile_header(), get_all_coords(), get_all_coords_vec(), massweights(), madness::plan_restart(), madness::MolecularOptimizer::projector_external_dof(), projector_external_dof(), test_ethylene(), and test_XCOperator_matrix().
◆ get_atom_charge()
| unsigned int madness::Molecule::get_atom_charge | ( | unsigned int | i | ) | const |
References atoms.
◆ get_atomic_number()
| unsigned int madness::Molecule::get_atomic_number | ( | unsigned int | i | ) | const |
References atoms.
Referenced by madness::DispersionCorrection::atomic_numbers(), madness::compare_geometry(), and get_structure().
◆ get_atoms()
|
inline |
References atoms.
Referenced by madness::Znemo::custom_guess().
◆ get_core_bc()
|
inline |
◆ get_core_l()
|
inline |
◆ get_eprec()
|
inline |
References parameters.
Referenced by madness::poly4erfc::poly4erfc(), madness::Slater::Slater(), add_atom(), add_atom(), read_core_file(), and update_rcut_with_eprec().
◆ get_pointgroup()
|
inline |
References pointgroup_.
◆ get_pseudo_atom()
| bool madness::Molecule::get_pseudo_atom | ( | unsigned int | i | ) | const |
References atoms.
Referenced by get_structure().
◆ get_rcut()
|
inline |
References rcut.
◆ get_structure()
| void madness::Molecule::get_structure | ( | ) |
References get_atomic_number(), madness::get_charge_from_file(), get_pseudo_atom(), MADNESS_EXCEPTION, natom(), orient(), parameters, read(), read_core_file(), read_structure_from_library(), read_xyz(), set_atom_charge(), and set_pseudo_atom().
Referenced by Molecule().
◆ get_structure_library_path()
|
static |
◆ guess_file()
|
inline |
References core_pot.
◆ hash()
|
inline |
References atoms, h(), madness::hash_combine(), madness::hash_range(), pointgroup_, and rcut.
◆ insert_symbols_and_geometry()
| void madness::Molecule::insert_symbols_and_geometry | ( | json & | mol_json | ) | const |
References madness::constants::atomic_unit_of_length, atoms, e(), madness::get_atomic_data(), natom(), parameters, and madness::scale().
Referenced by to_json(), and to_json_if_precedence().
◆ inter_atomic_distance()
| double madness::Molecule::inter_atomic_distance | ( | unsigned int | i, |
| unsigned int | j ) const |
References atoms, and madness::distance().
Referenced by nuclear_repulsion_derivative(), nuclear_repulsion_energy(), and nuclear_repulsion_second_derivative().
◆ is_potential_defined()
|
inline |
References core_pot.
◆ is_potential_defined_atom()
|
inline |
◆ massweights()
|
inline |
compute the mass-weighting matrix for the hessian
use as mass_weighted_hessian=inner(massweights,inner(hessian,massweights));
References get_atom(), and natom().
Referenced by madness::Nemo::compute_IR_intensities().
◆ molecular_core_potential()
| double madness::Molecule::molecular_core_potential | ( | double | x, |
| double | y, | ||
| double | z ) const |
References atoms, core_pot, madness::distance(), natom(), and madness::sum().
◆ moment_of_inertia()
| Tensor< double > madness::Molecule::moment_of_inertia | ( | ) | const |
References atoms, madness::FunctionDefaults< NDIM >::k, I, and q().
Referenced by madness::MolecularOptimizer::projector_external_dof(), and projector_external_dof().
◆ n_core_orb()
|
inline |
References core_pot.
◆ n_core_orb_all()
| unsigned int madness::Molecule::n_core_orb_all | ( | ) | const |
References atoms, core_pot, natom(), and madness::sum().
◆ natom()
|
inline |
References atoms.
Referenced by madness::DispersionCorrection::atomic_numbers(), center_of_mass(), madness::compare_geometry(), madness::DispersionCorrection::compute(), madness::Nemo::compute_IR_intensities(), core_potential_derivative(), cubefile_header(), get_all_coords(), get_all_coords_vec(), get_structure(), madness::DispersionCorrection::gradient(), madness::DispersionCorrection::hessian(), madness::Nemo::hessian(), insert_symbols_and_geometry(), madness::Nemo::make_incomplete_hessian(), madness::Nemo::make_incomplete_hessian_response_part(), massweights(), molecular_core_potential(), n_core_orb_all(), nuclear_repulsion_hessian(), madness::plan_restart(), print(), madness::MolecularOptimizer::projector_external_dof(), projector_external_dof(), read_core_file(), read_xyz(), set_all_coords(), madness::Nemo::solve_cphf(), test_gradient(), and test_XCOperator_matrix().
◆ nuclear_attraction_potential()
| double madness::Molecule::nuclear_attraction_potential | ( | double | x, |
| double | y, | ||
| double | z ) const |
nuclear attraction potential for the whole molecule
References atoms, madness::distance(), field, rcut, madness::smoothed_potential(), and madness::sum().
◆ nuclear_attraction_potential_derivative()
| double madness::Molecule::nuclear_attraction_potential_derivative | ( | int | atom, |
| int | axis, | ||
| double | x, | ||
| double | y, | ||
| double | z ) const |
References atoms, axis, madness::distance(), madness::dsmoothed_potential(), field, and rcut.
◆ nuclear_attraction_potential_second_derivative()
| double madness::Molecule::nuclear_attraction_potential_second_derivative | ( | int | atom, |
| int | iaxis, | ||
| int | jaxis, | ||
| double | x, | ||
| double | y, | ||
| double | z ) const |
the second derivative of the (smoothed) nuclear potential Z/r

with

References atoms, madness::d2smoothed_potential(), madness::Vector< T, N >::normf(), rcut, madness::smoothed_potential(), and u().
◆ nuclear_charge_density()
| double madness::Molecule::nuclear_charge_density | ( | double | x, |
| double | y, | ||
| double | z, | ||
| double | rscale = 1. ) const |
evaluate the nuclear charge density at point {x,y,z} using the default MADNESS nuclear model. See smoothed_density() for the description of the default nuclear model.
- Parameters
-
x,y,z the point at which the nuclear charge density is evaluated rscale setting rscale>1 will make a nucleus larger by a factor of rscale(in other words, rcut is multiplied by the inverse of by this)
- Returns
- the nuclear charge density at point {x,y,z}
- See also
- smoothed_density()
References atoms, madness::distance_sq(), MADNESS_ASSERT, rcut, and madness::smoothed_density().
◆ nuclear_dipole()
| double madness::Molecule::nuclear_dipole | ( | int | axis | ) | const |
compute the dipole moment of the nuclei
- Parameters
-
[in] axis the axis (x, y, z)
References atoms, axis, core_pot, MADNESS_EXCEPTION, and madness::sum().
◆ nuclear_dipole_derivative()
| Tensor< double > madness::Molecule::nuclear_dipole_derivative | ( | const int | atom, |
| const int | axis ) const |
compute the derivative of the nuclear dipole wrt a nuclear displacement
- Parameters
-
[in] atom the atom which will be displaced [in] axis the axis where the atom will be displaced
- Returns
- a vector which all 3 components of the dipole derivative
Referenced by madness::Nemo::compute_IR_intensities().
◆ nuclear_repulsion_derivative()
| double madness::Molecule::nuclear_repulsion_derivative | ( | size_t | iatom, |
| int | axis ) const |
References atoms, axis, core_pot, inter_atomic_distance(), and madness::sum().
◆ nuclear_repulsion_energy()
| double madness::Molecule::nuclear_repulsion_energy | ( | ) | const |
References atoms, core_pot, inter_atomic_distance(), and madness::sum().
◆ nuclear_repulsion_hessian()
| Tensor< double > madness::Molecule::nuclear_repulsion_hessian | ( | ) | const |
return the hessian matrix of the second derivatives d^2/dxdy V
compute the nuclear-nuclear contribution to the molecular hessian
no factor 0.5 included
References natom(), and nuclear_repulsion_second_derivative().
Referenced by madness::Nemo::hessian(), and madness::Nemo::make_incomplete_hessian().
◆ nuclear_repulsion_second_derivative()
| double madness::Molecule::nuclear_repulsion_second_derivative | ( | int | iatom, |
| int | jatom, | ||
| int | iaxis, | ||
| int | jaxis ) const |
compute the nuclear-nuclear contribution to the second derivatives
- Parameters
-
[in] iatom the i-th atom (row of the hessian) [in] jatom the j-th atom (column of the hessian) [in] iaxis the xyz axis of the i-th atom [in] jaxis the xyz axis of the j-th atom return the (3*iatom + iaxis, 3*jatom + jaxis) matix element of the hessian [in] iatom the i-th atom (row of the hessian) [in] jatom the j-th atom (column of the hessian) [in] iaxis the xyz axis of the i-th atom [in] jaxis the xyz axis of the j-th atom return the (3*iatom + iaxis, 3*jatom + jaxis) matix element
References atoms, core_pot, inter_atomic_distance(), MADNESS_EXCEPTION, and madness::sum().
Referenced by nuclear_repulsion_hessian().
◆ operator==()
|
inline |
References Molecule(), atoms, field, parameters, pointgroup_, and rcut.
◆ orient()
| void madness::Molecule::orient | ( | bool | verbose = false | ) |
Centers and orients the molecule in a standard manner.
References atoms, center(), e(), fix_phase(), madness::FunctionDefaults< NDIM >::k, I, MADNESS_CHECK, parameters, pointgroup_, madness::print(), q(), rotate(), madness::syev(), and symmetrize_and_identify_point_group().
Referenced by from_json(), and get_structure().
◆ position_stream_in_library()
|
static |
◆ print()
| void madness::Molecule::print | ( | std::ostream & | os = std::cout, |
| const bool | defined_only = false ) const |
print all parameters and the molecular geometry
References atoms, madness::get_atomic_data(), natom(), p(), parameters, and madness::AtomicData::symbol.
Referenced by madness::SCF::load_mos(), main(), print_defined_only(), and test_ethylene().
◆ print_defined_only()
|
inline |
print user-defined parameters and the molecular geometry
References print().
◆ print_parameters()
|
static |
References param, and madness::print().
Referenced by madness::CC2::print_parameters(), madness::MP2::print_parameters(), madness::Nemo::print_parameters(), madness::OEP::print_parameters(), madness::PNO::print_parameters(), madness::SCF::print_parameters(), madness::TDHF::print_parameters(), madness::Zcis::print_parameters(), and madness::Znemo::print_parameters().
◆ read()
| void madness::Molecule::read | ( | std::istream & | f | ) |
References add_atom(), madness::constants::atomic_unit_of_length, atoms, madness::check_if_pseudo_atom(), e(), madness::f, p(), parameters, madness::position_stream_at_coordinates(), rcut, madness::scale(), madness::symbol_to_atomic_number(), tag, and update_rcut_with_eprec().
Referenced by get_structure(), read_file(), and read_structure_from_library().
◆ read_core_file()
| void madness::Molecule::read_core_file | ( | const std::string & | filename | ) |
References atoms, core_pot, madness::filename, get_eprec(), natom(), madness::print(), q(), rcut, and madness::smoothing_parameter().
Referenced by Molecule(), and get_structure().
◆ read_file()
| void madness::Molecule::read_file | ( | const std::string & | filename | ) |
References errmsg(), madness::f, madness::filename, MADNESS_EXCEPTION, and read().
◆ read_structure_from_library()
| void madness::Molecule::read_structure_from_library | ( | const std::string & | name | ) |
References errmsg(), madness::f, MADNESS_EXCEPTION, madness::name(), position_stream_in_library(), and read().
Referenced by get_structure().
◆ read_xyz()
| void madness::Molecule::read_xyz | ( | const std::string | filename | ) |
References add_atom(), madness::constants::atomic_unit_of_length, atoms, madness::check_if_pseudo_atom(), e(), errmsg(), madness::f, madness::filename, MADNESS_CHECK, MADNESS_EXCEPTION, natom(), parameters, rcut, madness::scale(), madness::symbol_to_atomic_number(), tag, and update_rcut_with_eprec().
Referenced by get_structure().
◆ rotate()
| void madness::Molecule::rotate | ( | const Tensor< double > & | D | ) |
- Parameters
-
[in] D the rotation matrix
References atoms, field, and madness::inner().
Referenced by orient().
◆ serialize()
|
inline |
References atoms, core_pot, field, parameters, pointgroup_, and rcut.
◆ set_all_coords()
| void madness::Molecule::set_all_coords | ( | const madness::Tensor< double > & | newcoords | ) |
References atoms, c, MADNESS_ASSERT, and natom().
Referenced by test_gradient().
◆ set_atom_charge()
| void madness::Molecule::set_atom_charge | ( | unsigned int | i, |
| double | zeff ) |
References atoms.
Referenced by get_structure().
◆ set_atom_coords()
| void madness::Molecule::set_atom_coords | ( | unsigned int | i, |
| double | x, | ||
| double | y, | ||
| double | z ) |
References atoms.
◆ set_core_eprec()
|
inline |
References core_pot, and madness::Future< void >::value.
◆ set_core_rcut()
|
inline |
References core_pot, and madness::Future< void >::value.
◆ set_pseudo_atom()
| void madness::Molecule::set_pseudo_atom | ( | unsigned int | i, |
| bool | psat ) |
References atoms.
Referenced by get_structure().
◆ set_rcut()
| void madness::Molecule::set_rcut | ( | double | value | ) |
References atoms, madness::Future< void >::value, and rcut.
◆ smallest_length_scale()
◆ swapaxes()
|
private |
Referenced by symmetrize_and_identify_point_group().
◆ symmetrize_and_identify_point_group()
| std::string madness::Molecule::symmetrize_and_identify_point_group | ( | const double | symtol | ) |
References madness::inverse(), madness::print(), swapaxes(), symmetrize_for_op(), test_for_c2(), test_for_inverse(), and test_for_sigma().
Referenced by orient().
◆ symmetrize_for_op()
|
private |
References atoms, madness::distance(), find_symmetry_equivalent_atom(), MADNESS_CHECK, op(), and test_for_op().
Referenced by symmetrize_and_identify_point_group().
◆ test_for_c2()
|
private |
References test_for_op().
Referenced by symmetrize_and_identify_point_group().
◆ test_for_inverse()
|
private |
References test_for_op().
Referenced by symmetrize_and_identify_point_group().
◆ test_for_op()
|
private |
References atoms, find_symmetry_equivalent_atom(), and op().
Referenced by symmetrize_for_op(), test_for_c2(), test_for_inverse(), and test_for_sigma().
◆ test_for_sigma()
|
private |
References test_for_op().
Referenced by symmetrize_and_identify_point_group().
◆ to_json()
|
nodiscard |
References insert_symbols_and_geometry(), and parameters.
Referenced by madness::MolecularEnergy::output_calc_info_schema(), and to_json_if_precedence().
◆ to_json_if_precedence()
|
inlinenodiscard |
References insert_symbols_and_geometry(), parameters, and to_json().
◆ total_nuclear_charge()
| double madness::Molecule::total_nuclear_charge | ( | ) | const |
References atoms, and madness::sum().
◆ translate()
| void madness::Molecule::translate | ( | const Tensor< double > & | translation | ) |
translate the molecule
References atoms.
Referenced by center(), madness::MolecularOptimizer::projector_external_dof(), and projector_external_dof().
◆ update_rcut_with_eprec()
| void madness::Molecule::update_rcut_with_eprec | ( | double | value | ) |
updates rcuts with given eprec
References atoms, core_pot, madness::Future< void >::value, get_eprec(), parameters, rcut, and madness::smoothing_parameter().
Referenced by dnuclear_anchor_test(), read(), and read_xyz().
Member Data Documentation
◆ atomic_radii
| std::vector<double> madness::Molecule::atomic_radii |
Referenced by Molecule(), add_atom(), and add_atom().
◆ atoms
|
private |
Referenced by Molecule(), Molecule(), Molecule(), add_atom(), add_atom(), atomic_attraction_potential(), bounding_cube(), center(), core_derivative(), core_eval(), core_potential_derivative(), find_symmetry_equivalent_atom(), fix_phase(), from_json(), get_atom(), get_atom_charge(), get_atomic_number(), get_atoms(), get_pseudo_atom(), hash(), insert_symbols_and_geometry(), inter_atomic_distance(), is_potential_defined_atom(), molecular_core_potential(), moment_of_inertia(), n_core_orb_all(), natom(), nuclear_attraction_potential(), nuclear_attraction_potential_derivative(), nuclear_attraction_potential_second_derivative(), nuclear_charge_density(), nuclear_dipole(), nuclear_dipole_derivative(), nuclear_repulsion_derivative(), nuclear_repulsion_energy(), nuclear_repulsion_second_derivative(), operator==(), orient(), print(), read(), read_core_file(), read_xyz(), rotate(), serialize(), set_all_coords(), set_atom_charge(), set_atom_coords(), set_pseudo_atom(), set_rcut(), smallest_length_scale(), swapaxes(), symmetrize_for_op(), test_for_op(), total_nuclear_charge(), translate(), and update_rcut_with_eprec().
◆ core_pot
|
private |
Referenced by Molecule(), Molecule(), Molecule(), core_derivative(), core_eval(), core_potential_derivative(), get_core_bc(), get_core_l(), guess_file(), is_potential_defined(), is_potential_defined_atom(), molecular_core_potential(), n_core_orb(), n_core_orb_all(), nuclear_dipole(), nuclear_repulsion_derivative(), nuclear_repulsion_energy(), nuclear_repulsion_second_derivative(), read_core_file(), serialize(), set_core_eprec(), set_core_rcut(), and update_rcut_with_eprec().
◆ field
|
private |
◆ parameters
| GeometryParameters madness::Molecule::parameters |
◆ pointgroup_
|
private |
The molecular point group is automatically assigned in the identify_pointgroup function
Referenced by get_pointgroup(), hash(), operator==(), orient(), and serialize().
◆ rcut
|
private |
Referenced by Molecule(), Molecule(), add_atom(), add_atom(), atomic_attraction_potential(), from_json(), get_rcut(), hash(), nuclear_attraction_potential(), nuclear_attraction_potential_derivative(), nuclear_attraction_potential_second_derivative(), nuclear_charge_density(), operator==(), read(), read_core_file(), read_xyz(), serialize(), set_rcut(), smallest_length_scale(), and update_rcut_with_eprec().
◆ tag
|
staticconstexpr |
The documentation for this class was generated from the following files:
Generated on for MADNESS by
 1.15.0