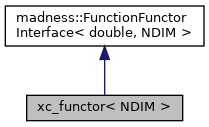
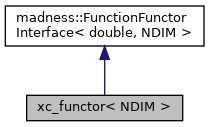
Functor for the exchange correlation potential. More...
Public Member Functions | |
| xc_functor () | |
| xc_functor (std::vector< atom_information< NDIM > > atoms) | |
| double | operator() (const Vector< double, NDIM > &r) const |
| returns the slater potential | |
| Public Member Functions inherited from madness::FunctionFunctorInterface< double, NDIM > | |
| FunctionFunctorInterface () | |
| virtual | ~FunctionFunctorInterface () |
| virtual coeffT | coeff (const keyT &) const |
| virtual void | operator() (const Vector< double *, 1 > &, double *, int) const |
| virtual bool | provides_coeff () const |
| does this functor directly provide sum coefficients? or only function values? | |
| virtual bool | screened (const Vector< double, NDIM > &, const Vector< double, NDIM > &) const |
| Can we screen this function based on the bounding box information? | |
| void | set_length_scale (double lo) |
| adapt the special level to resolve the smallest length scale | |
| virtual Level | special_level () const |
| Override this to change the minimum level of refinement at special points (default is 6). | |
| virtual std::vector< Vector< double, NDIM > > | special_points () const |
| Override this to return list of special points to be refined more deeply. | |
| virtual bool | supports_vectorized () const |
| Does the interface support a vectorized operator()? | |
| virtual coeffT | values (const keyT &, const Tensor< double > &) const |
Private Member Functions | |
| double | density (const Vector< double, NDIM > &r) const |
| forms the density | |
| double | get_distance (Vector< double, NDIM > elec, Vector< double, NDIM > nuc) const |
| computes distance between coordinate of electron and coordinate of nucleus | |
| double | xc_potential (const Vector< double, NDIM > &r) const |
| equation for the slater potential | |
Private Attributes | |
| std::vector< atom_information< NDIM > > | atoms |
| Needed information about the molecule to apply asymtotic correction. | |
Additional Inherited Members | |
| Public Types inherited from madness::FunctionFunctorInterface< double, NDIM > | |
| typedef GenTensor< double > | coeffT |
| typedef Key< NDIM > | keyT |
| typedef double | value_type |
| Public Attributes inherited from madness::FunctionFunctorInterface< double, NDIM > | |
| Level | special_level_ |
Detailed Description
class xc_functor< NDIM >
Functor for the exchange correlation potential.
Asymptotic correction for DFT. In the correction the xc-potential is replaced by an 1/r term far away from the nuclei to give the correct asymptotic behavior. Close to the nuclei the standard xc-potential is used. The transition between the different potentials is achieved via a linear interpolation. This is a test code to compute the corrected potential in 1D or 2D. The density functional is computed using the slater potential. The orbitals have to be inserted manually in the xc_functor class in the density() function. The molecule/atom has to be initialized manually in the main function and has to be stored as a vector of atom_information.
Constructor & Destructor Documentation
◆ xc_functor() [1/2]
|
inline |
◆ xc_functor() [2/2]
|
inline |
References atoms.
Member Function Documentation
◆ density()
|
inlineprivate |
forms the density
References atoms, density(), and get_distance().
Referenced by density(), and xc_potential().
◆ get_distance()
|
inlineprivate |
computes distance between coordinate of electron and coordinate of nucleus
References madness::distance(), and NDIM.
Referenced by density().
◆ operator()()
|
inline |
returns the slater potential
References xc_potential().
◆ xc_potential()
|
inlineprivate |
Member Data Documentation
◆ atoms
|
private |
Needed information about the molecule to apply asymtotic correction.
Referenced by xc_functor(), and density().
The documentation for this class was generated from the following file:
Generated on for MADNESS by
 1.15.0